
Unmapped Read From Bam Files Issue 1153 Samtoolssamtools Github
unmapped read from bam files · Issue #1153 · samtools/samtools · GitHub
Multithreaded reading of BAM file · Issue #235 · samtools/samtools · GitHub
Are Unmapped reads in the BAM file corrected ? or discarded ? · Issue ...
convert bam to fastq for each read group in bam file · Issue #1032 ...
GitHub - anands-repo/bamutils: Some tools for manipulating BAM files ...
Extract Unmapped Reads : Extract paired FASTQ reads from a BAM mapping ...
mpileup on multiple files only works if they are bam · Issue #123 ...
samtools header read error from empty file · Issue #191 · STAR-Fusion ...

Extract Unmapped Reads : Extract paired FASTQ reads from a BAM mapping ...
Unable to read file error when using long list of Bam files - solution ...
Advertisement Space (300x250)
Trouble reading SRA CRAM files · Issue #1254 · samtools/samtools · GitHub
GitHub - Sydney-Informatics-Hub/Fix-BAM-read-groups: Change the read ...
GitHub - jmschrei/bam2bw: A command-line tool for reading SAM/BAM files ...

Missing mpileup records · Issue #1311 · samtools/bcftools · GitHub
`samtools index` silently fails to detect corrupted BAM files and ...
Convert BAM file to fast file - Error · Issue #1344 · samtools/samtools ...
Samtools index fails on a sorted bam file · Issue #1126 · samtools ...
`--input-fmt-option` filter not work on unmapped reads · Issue #1445 ...
undefined reference to 'bam_dup1' · Issue #192 · samtools/samtools · GitHub
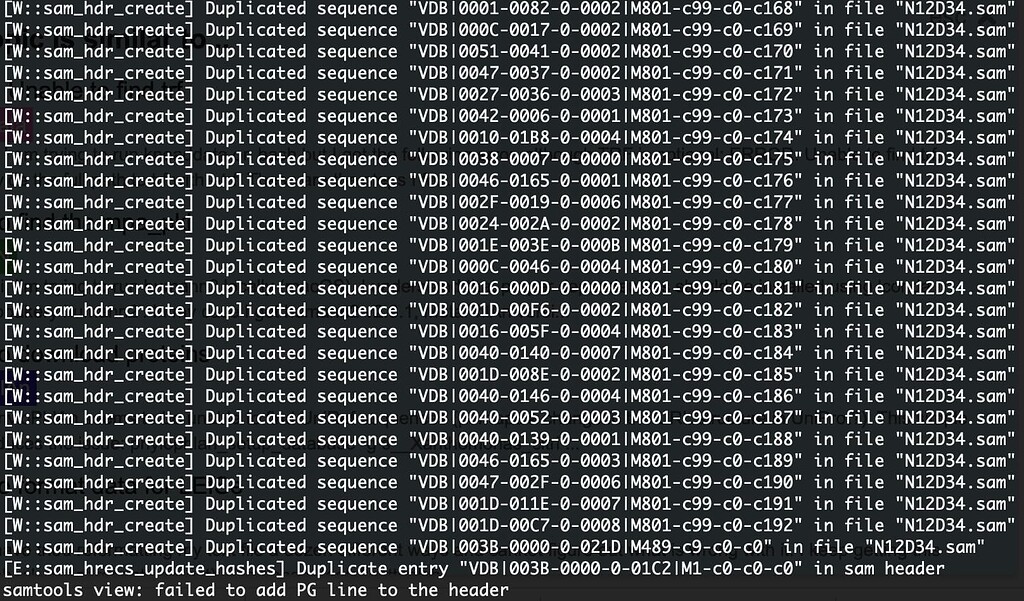
Samtools failed to read header of sam file · Issue #1774 · samtools ...
Advertisement Space (336x280)
Error encountered while indexing BAM file · Issue #1559 · samtools ...
View First Few Records of BAM File · Issue #943 · samtools/samtools ...
Error "Failed to read from standard input: unknown file type" with ...
Default behavior - save unmapped reads to SAM/BAM output · Issue #44 ...
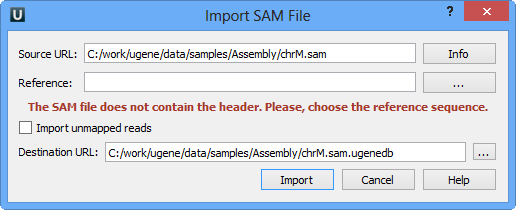
Import BAM and SAM Files | UGENE Documentation
samtools error ([main_samview] fail to read the header from "-") during ...
Incorrect BAM? · Issue #1111 · samtools/samtools · GitHub
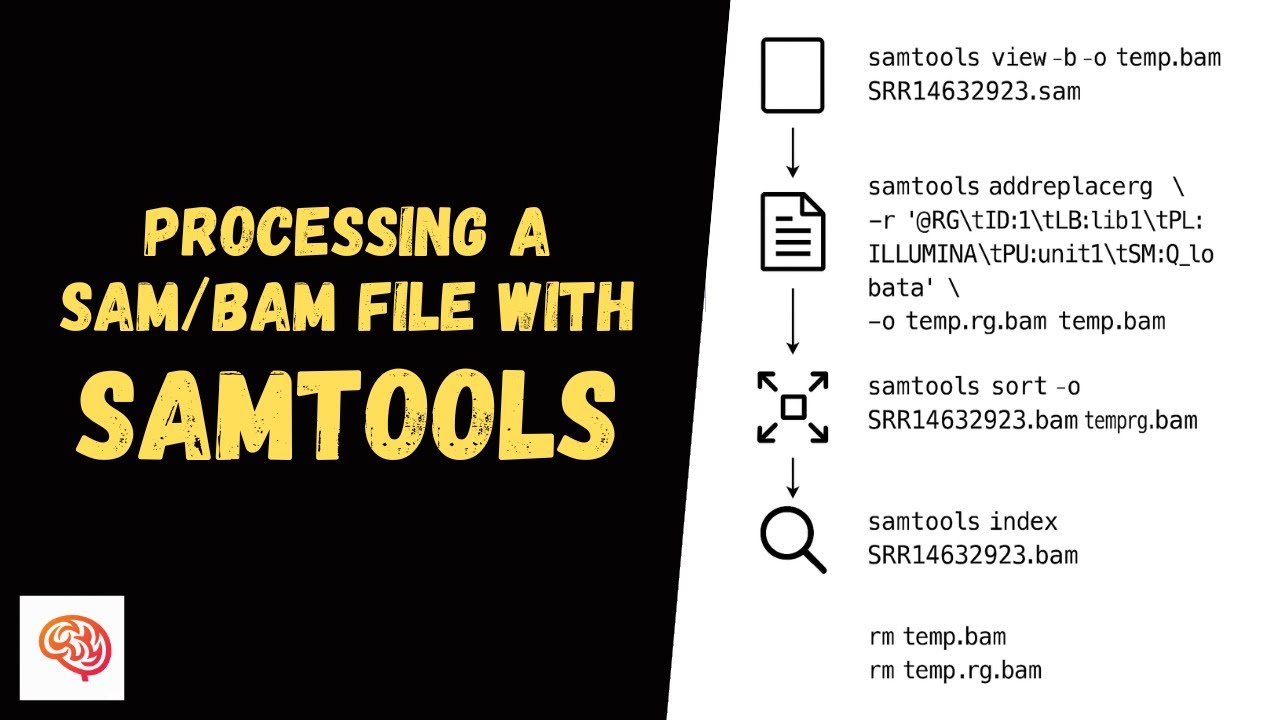
How to Convert, Add Read Group, Sort and Index SAM/BAM Files with ...
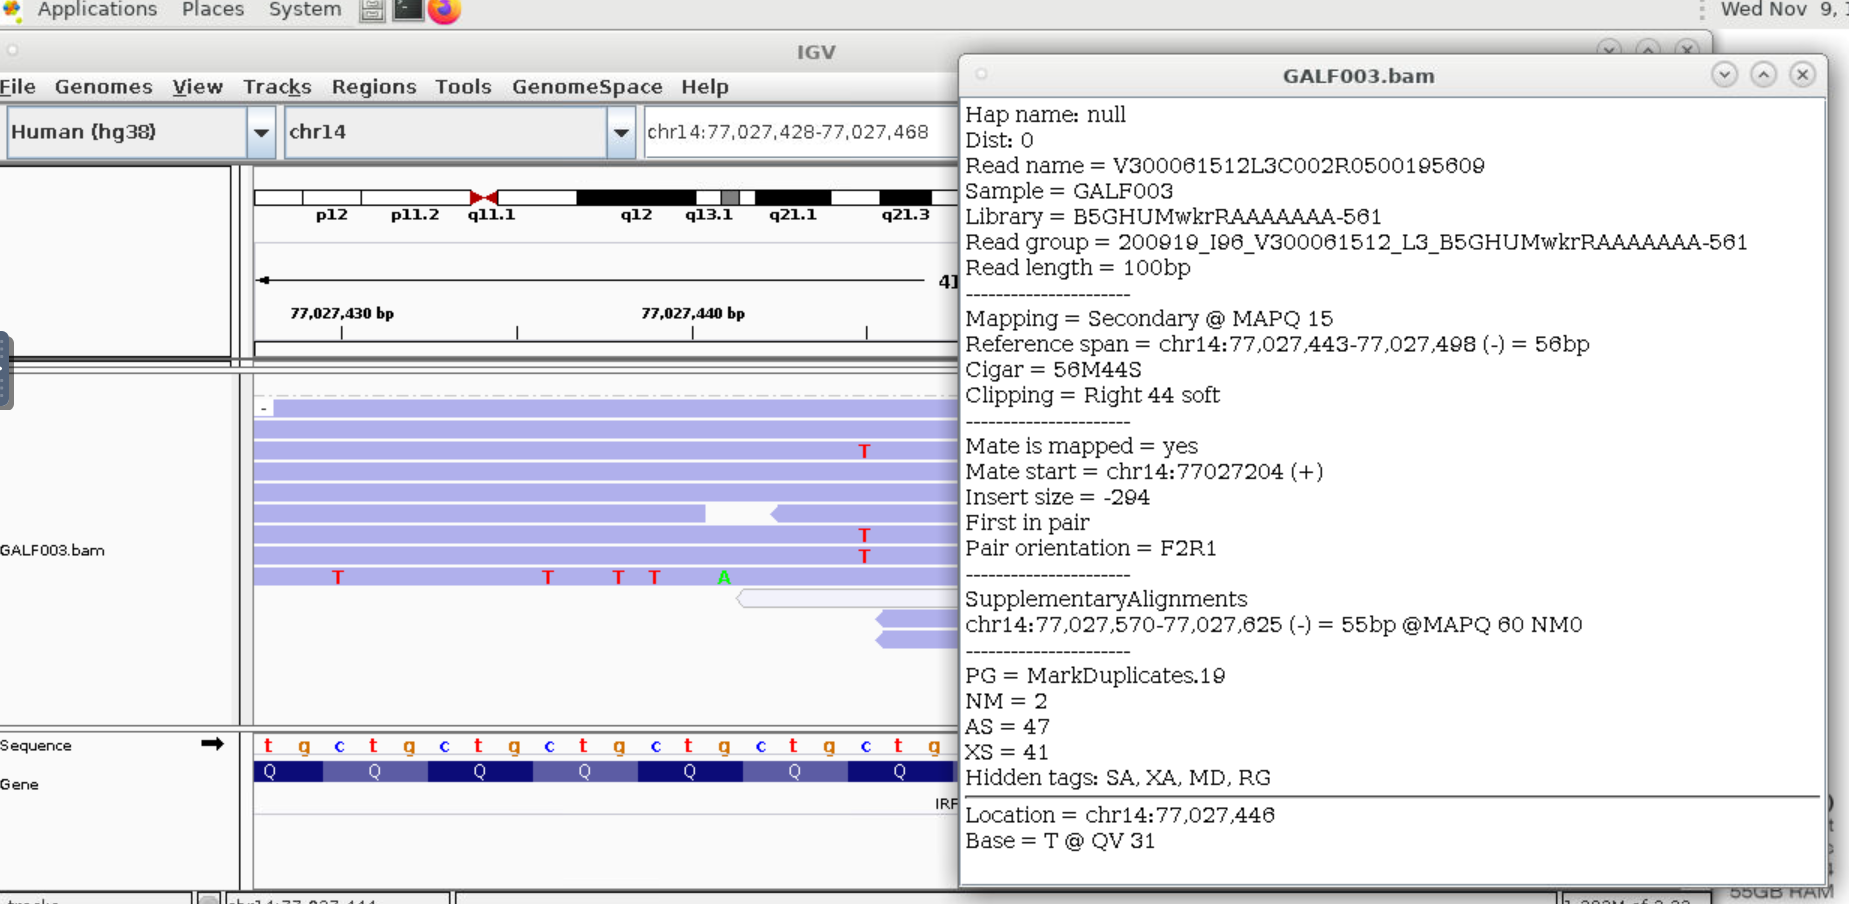
Secondary read missing when compared with IGV · Issue #1753 · samtools ...

Confused about % of mapped and unmapped reads output from STAR aligner
Advertisement Space (336x280)
Shift reads in sam/bam files directly · Issue #868 · samtools/samtools ...

Samtools mpileup information · Issue #1282 · samtools/samtools · GitHub
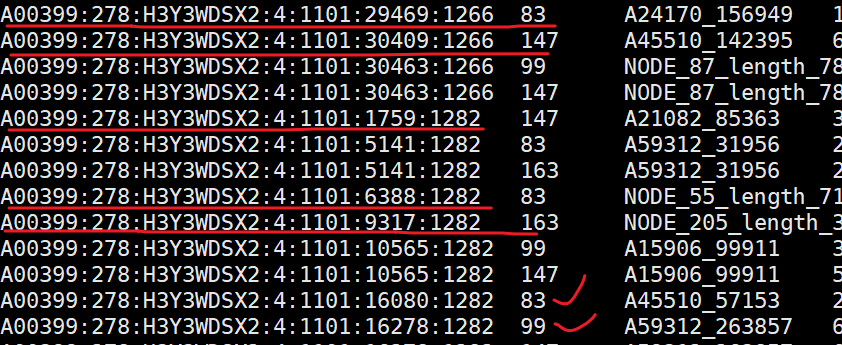
samtools filter by single reads · Issue #1768 · samtools/samtools · GitHub
Bam2fq error · Issue #1892 · samtools/samtools · GitHub
can not get the bam files using pipeline bwa mem | samtools view -Sb ...
Fail to index using samtools for a bam file · Issue #1141 · samtools ...